An EU MDR audit is a rigorous conformity assessment performed by a Notified Body to verify that a medical device manufacturer’s Quality Management System (QMS) and technical documentation comply with Regulation (EU) 2017/745. This assessment is mandatory for placing or maintaining medical devices on the European Union market, ensuring clinical safety and performance.
Understanding the EU MDR Conformity Assessment
The European Union Medical Device Regulation (EU MDR 2017/745) shifted the landscape of device compliance from a reactive, certificate-based model to a continuous, lifecycle-oriented oversight. An EU MDR audit is not a one-time administrative hurdle; it is a systematic examination of a manufacturer’s processes, design data, and clinical evidence.
The primary objective of these audits is to verify that a manufacturer maintains a robust QMS—typically aligned with ISO 13485:2016—while ensuring that the technical documentation for each device (or device family) meets the high standards of the regulation. Because the EU does not "approve" devices in the same way the FDA does in the United States, the audit serves as the gateway for a Notified Body to issue a CE Certificate, which permits the manufacturer to affix the CE mark and circulate the product throughout the European Economic Area.
Prerequisites for a Successful Audit
Before scheduling an audit, a manufacturer must ensure they have established the foundational infrastructure required by the regulation. Attempting to bypass these prerequisites often leads to "Major Non-Conformities," which can halt the certification process and jeopardize market access.
- QMS Implementation: Your Quality Management System must be documented, implemented, and maintained. It should cover the entire product lifecycle, from design and development to post-market surveillance.
- Technical Documentation: Each device must have a complete technical file. This includes the General Safety and Performance Requirements (GSPR), risk management files, clinical evaluation reports (CER), and labeling/instructions for use.
- Unique Device Identification (UDI): Ensure your devices are registered in the EUDAMED database and correctly labeled with the appropriate UDI-DI and UDI-PI codes.
- Person Responsible for Regulatory Compliance (PRRC): Per Article 15 of the MDR, you must have a designated individual within your organization who possesses the necessary expertise to ensure regulatory compliance.
- Notified Body Designation: You must engage a designated Notified Body (NB) with the specific "scope" of certification that matches your device’s classification and technology type.
The Stages of the EU MDR Audit
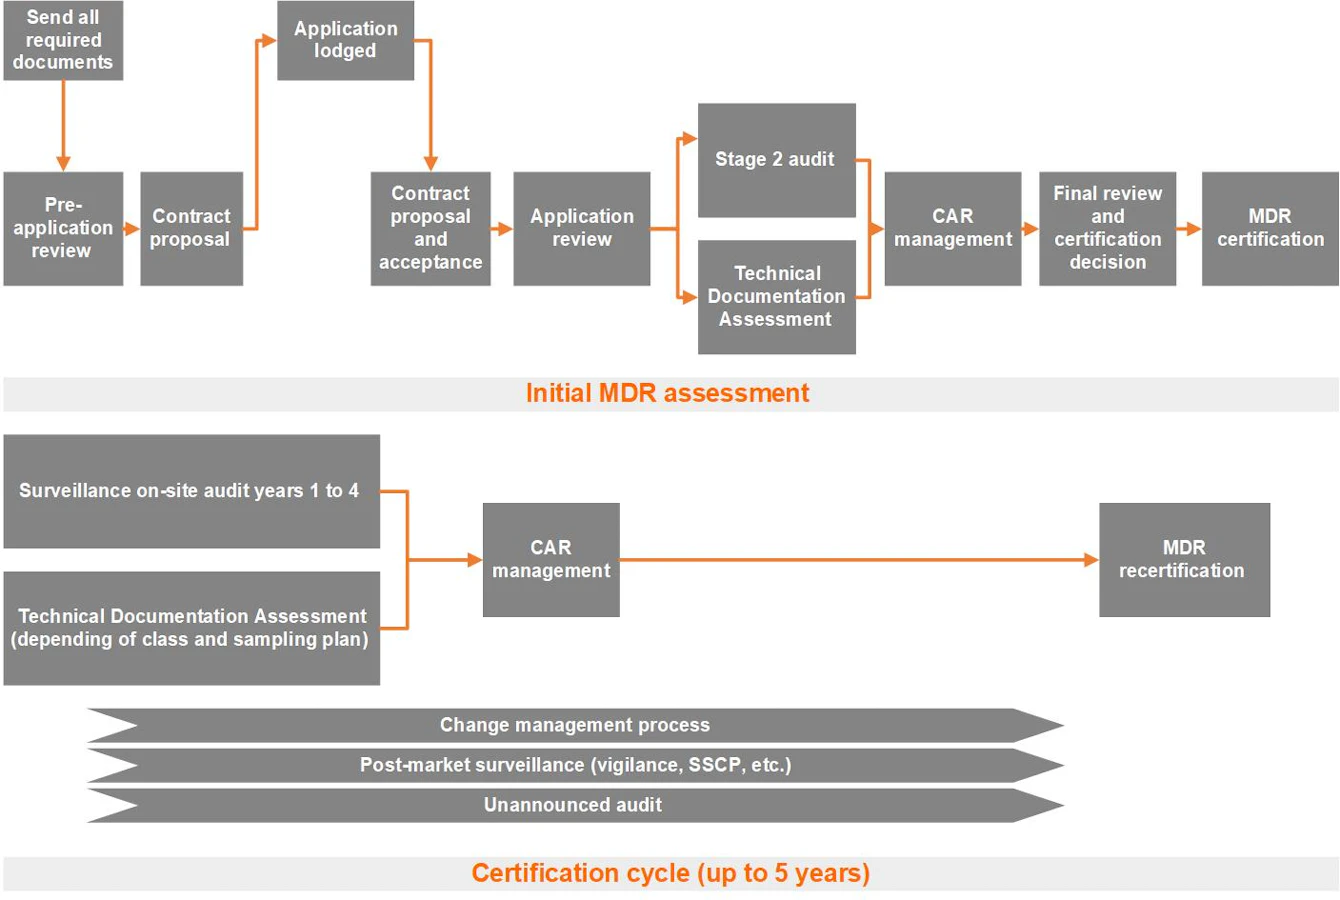
The audit process is structured into two main phases, often referred to as Stage 1 and Stage 2 audits, followed by ongoing surveillance.
Stage 1: Document Review
The purpose of the Stage 1 audit is to determine the manufacturer’s readiness for the full-scale Stage 2 assessment. The Notified Body auditors review your documentation to ensure that your QMS policies are compliant on paper. They will verify that you have identified the applicable GSPR for your devices and that your clinical evaluation strategy is logical and grounded in evidence. If the auditors find significant gaps at this stage, they will provide a report, allowing the manufacturer to rectify issues before the onsite audit.
Stage 2: System and Product Verification
This is the core of the audit. Auditors verify the implementation of the processes reviewed in Stage 1. They will conduct interviews with staff, observe production lines, inspect cleanrooms, and perform a deep dive into the technical documentation of sample devices. The goal is to prove that the QMS is not just a collection of manuals, but a functional framework that consistently produces safe and effective devices.
Surveillance Audits
Under the EU MDR, certification cycles are shorter and more intense than under the previous Medical Device Directive (MDD). You can expect annual surveillance audits, along with unannounced audits. Unannounced audits have become a standard feature of the MDR, intended to ensure that manufacturing consistency is maintained even when auditors are not scheduled to visit.
Managing Non-Conformities
It is rare for an audit to conclude with zero findings. Auditors categorize findings into two levels:
- Minor Non-Conformity: A lapse in compliance that does not fundamentally undermine the QMS or the safety of the device. These must be addressed through a formal Corrective Action plan by a specified deadline.
- Major Non-Conformity: A systemic failure in the QMS or a lack of objective evidence for safety/performance. A major non-conformity will prevent the issuance of a CE certificate until the auditor verifies that effective corrective measures have been implemented.
Comparison of Audit Focus Areas
| Audit Focus | Primary Objective | Key Evidence Required |
|---|---|---|
| QMS Audit | Verify system effectiveness | SOPs, internal audit reports, management reviews |
| Technical File | Verify product safety | GSPR checklist, risk management files, test reports |
| Clinical Evaluation | Verify clinical performance | CER, clinical investigation data, literature reviews |
| Post-Market (PMS) | Verify real-world performance | PMS plan, PSUR (Periodic Safety Update Report) |
Common Pitfalls and How to Avoid Them
Many manufacturers stumble when transitioning from MDD to MDR, or when entering the EU market for the first time. The most frequent errors involve:
- Insufficient Clinical Data: The MDR requires "clinical sufficiency." Relying solely on equivalence to a competitor’s device is no longer sufficient; you must often provide direct clinical evidence for your specific device.
- Inadequate Risk Management: Risk management must be a continuous process, not a pre-market task. Auditors will look for evidence that you are updating your risk assessments based on real-world incident data.
- Scope Mismatch: Many delays occur because the manufacturer’s chosen Notified Body does not have the specific technological scope (e.g., software-as-a-medical-device or active implants) to certify the product.
- Language and Translation Errors: Technical documentation must be accurate and available in the languages required by the member states where you intend to sell, particularly for labeling and user instructions.
The Role of EUDAMED
By 2026, the European Database on Medical Devices (EUDAMED) remains a central component of the audit process. While some modules are mandatory and others are gradually being rolled out, manufacturers are expected to maintain current, accurate data within the portal. Auditors will cross-reference the data present in your technical documentation with the information registered in EUDAMED. Discrepancies between your internal files and the central database are considered a significant compliance risk.
Clinical Evaluation Reports (CER)
The CER is the cornerstone of your audit defense. Under the EU MDR, your clinical evaluation must be updated regularly. If your device is a high-risk implantable, you may be required to conduct a Post-Market Clinical Follow-up (PMCF) study. Auditors will scrutinize the scientific validity of the data you present. If your clinical evidence is deemed insufficient, the audit will likely result in a rejection or a requirement for further, potentially costly, clinical investigations.
Professional Advice and Liability
This guide provides an overview of the regulatory landscape as of 2026. The EU MDR is a complex legal framework, and the interpretation of these requirements can vary between different Notified Bodies and EU member states. Manufacturers are strongly encouraged to consult with a qualified regulatory affairs consultant or legal counsel specialized in European medical device law. Compliance is ultimately the legal responsibility of the manufacturer; outsourcing activities to consultants does not transfer the ultimate liability for device safety or regulatory failure.
Frequently Asked Questions
How long does an EU MDR audit take? The duration varies based on the size of the manufacturer and the complexity of the device portfolio. A typical audit process, including document review and onsite inspection, usually spans several months, with the certification process taking anywhere from 9 to 18 months depending on the responsiveness of the manufacturer and the current capacity of the Notified Body.
What happens if I fail an audit? You do not receive a certificate, and you cannot place or continue to sell your devices on the EU market. You must perform a root cause analysis, implement corrective actions, and submit to a follow-up assessment. The length of time required to rectify issues depends entirely on the nature of the non-conformities found.
Are unannounced audits mandatory? Yes, under the EU MDR, Notified Bodies are required to conduct unannounced audits at least once every three years. These are designed to ensure that the production processes you presented during your initial audit are being followed every single day.
Does an FDA clearance help with an EU MDR audit? While having FDA approval (510(k) or PMA) demonstrates a high level of rigor and maturity, it does not directly equate to MDR compliance. The documentation requirements are different, and the EU MDR often requires more exhaustive clinical data. Use your FDA documentation as a foundation, but expect to perform a significant gap analysis to align it with MDR requirements.
Next Steps for Manufacturers
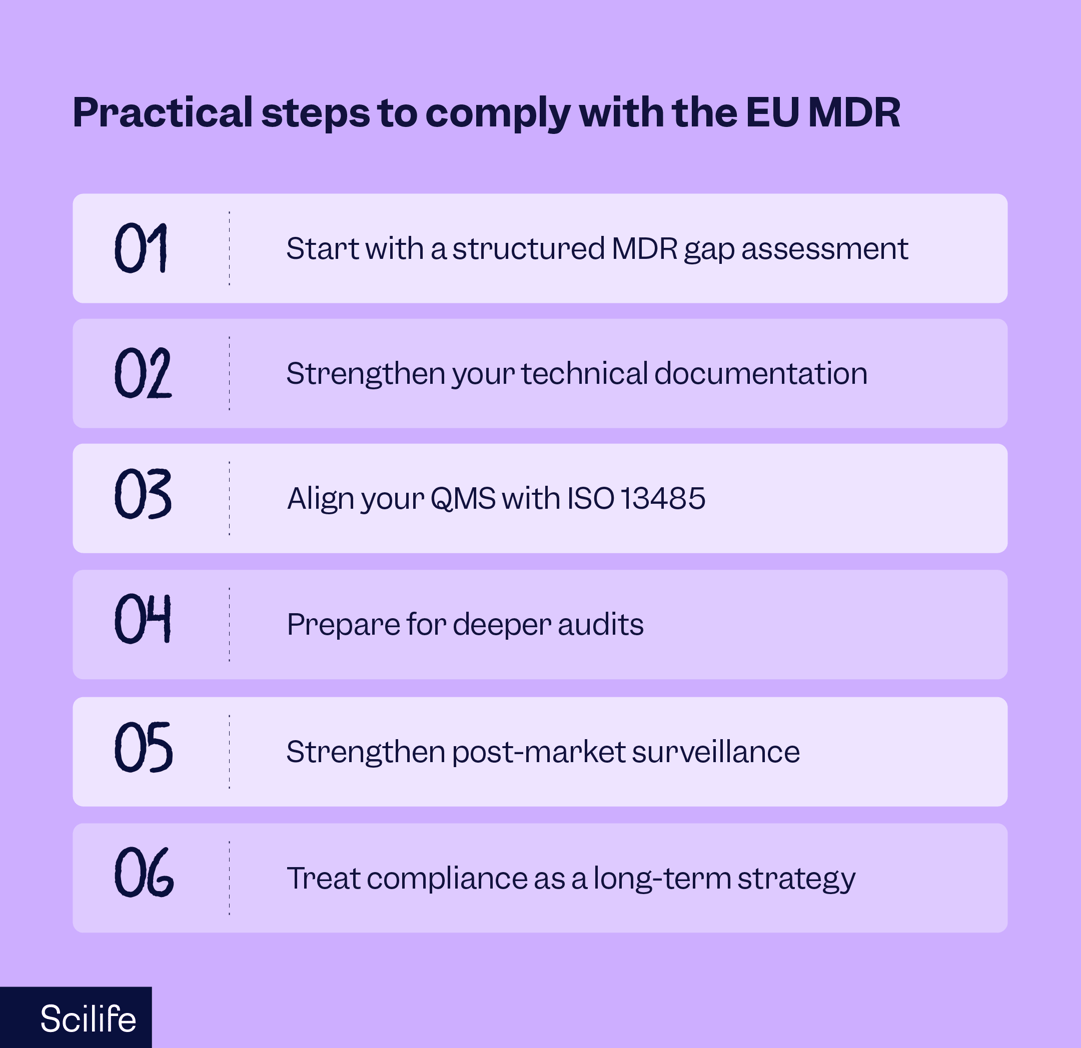
If you are preparing for an upcoming audit, the first step is to conduct a formal gap analysis against the current MDR requirements. Review your technical documentation for consistency, ensure your UDI entries in EUDAMED are up to date, and perform an internal audit of your QMS. Engage your Notified Body early, maintain clear communication, and ensure your PRRC is actively involved in every phase of the preparation. Success in the EU market depends on proactive vigilance, not just reactive compliance.